Historical profile. Long before anyone could sequence a gene for appetite, Douglas Coleman did something that sounds crude today.
At The Jackson Laboratory in Bar Harbor, Maine, he surgically joined pairs of mice so that they shared a circulation, then watched how their feeding behavior, weight and blood sugar changed.
Some of the animals stopped eating and starved. Others gained weight more slowly. The outcome depended on which genetically obese and normal mice were paired.
From those experiments, Douglas Coleman concluded that blood carried a signal involved in controlling hunger and body fat, and that two strains of obese mice had defects at different points in the same biological pathway.
He could not identify the molecule. But the experiments allowed him to predict that one mouse strain lacked a circulating signal while the other produced it but could not respond.
More than two decades later, in 1994, researchers identified the hormone and named it leptin.
This is the story of how a biochemist working at a mouse genetics laboratory helped establish that obesity could be regulated by hormones, long before modern molecular biology could explain exactly how.
A Biochemist in a Mouse Genetics Laboratory
Douglas L. Coleman was born in 1931 in Stratford, Ontario, Canada.
He was an only child and grew up in modest circumstances. His son, Thomas R. Coleman, later described a household where his father developed an early fascination with understanding how things worked.
Coleman studied at McMaster University in Hamilton, Ontario, where he met his future wife, Beverly Benallick.
He continued his education at the University of Wisconsin–Madison, studying biochemistry under Carl Baumann, and completed his PhD in 1958.
That same year, he joined The Jackson Laboratory in Bar Harbor, Maine, an institution known for developing and maintaining genetically defined mouse models used in biomedical research.
He initially expected to remain there for only a short period. Instead, he spent his entire scientific career at the laboratory.
One of the institution’s most unusual research resources was a strain of mice carrying a mutation called ob, first described in a 1950 publication.
Mice inheriting two copies of this mutation, known as ob/ob mice, developed extreme obesity, excessive appetite and abnormalities in glucose metabolism.
In 1965, researchers discovered another spontaneous mutation that produced a similar-looking animal.
This second mutation, named db for diabetes, was reported in a 1966 Science paper authored by Katherine Hummel, Margaret Dickie and Coleman.
Both mutant strains became severely obese, but the db/db animals also developed particularly serious diabetes on the genetic background in which they were originally studied.
Coleman and his colleagues demonstrated that genetic background could substantially change the severity of diabetes associated with these mutations.
That finding suggested that obesity and diabetes were related conditions, but that their severity was not determined by a single mutation acting in isolation.
The next question was more fundamental: why did two different mutations produce animals that appeared so similar?
Douglas Coleman and the Parabiosis Experiments
Parabiosis is an experimental technique in which two animals are surgically joined so that a shared blood circulation develops between them.
It was not a new invention. Earlier researchers, including G. R. Hervey in 1959, had used parabiosis to investigate possible circulating signals involved in body-weight regulation.
Coleman’s contribution was to use the technique to compare two different genetic causes of extreme obesity.
The reasoning was straightforward.
If obesity resulted from the absence of something normally found in blood, attaching an affected animal to a healthy one might supply the missing substance.
If a genetically obese animal produced an unusually powerful circulating substance, that substance might affect its partner.
The first major test paired db/db mice with normal mice.
In the experiments published by Coleman and Katherine Hummel in the American Journal of Physiology in 1969, the normal partners developed severe metabolic abnormalities while the genetically diabetic animals continued eating and gaining weight.
The normal mice eventually died.
Coleman recalled that he initially worried the deaths reflected a failure of his surgical technique.
But the outcome happened repeatedly, and the evidence pointed toward starvation rather than an ordinary surgical complication.
After approximately one week, blood glucose in the normal partners had fallen to starvation levels.
Postmortem examinations found little or no food in their digestive tracts and no detectable glycogen stores in their livers.
The db/db partners, meanwhile, remained hyperglycaemic and continued consuming food.
Coleman interpreted the result as evidence that the genetically diabetic mouse produced a powerful circulating factor capable of suppressing appetite in its normal partner.
The normal mouse starved while the obese partner kept eating. Coleman suspected the difference was not appetite alone, but an invisible signal carried through their shared blood.
That was a remarkable observation. It suggested that an animal could become obese not because it lacked a signal to stop eating, but because it could no longer respond to that signal.
Two Obese Mouse Mutations, Two Different Defects
The next challenge was to compare the two mutant strains directly.
Initially, the ob/ob and db/db mice came from different genetic backgrounds, creating a risk of immune rejection when the animals were surgically joined.
Coleman used breeding strategies to place the mutations onto compatible backgrounds, allowing the crucial parabiosis comparisons to proceed.
The findings were reported in a 1973 paper in Diabetologia.
When an ob/ob mouse was joined to a db/db mouse, the outcome resembled the earlier experiment with a normal partner.
The db/db mouse continued feeding and gaining weight. The ob/ob partner reduced its food intake, lost substantial body fat, developed very low blood glucose and eventually died of starvation.
In his later account, Coleman described survival periods of approximately 20 to 30 days for these ob/ob partners.
But when an ob/ob mouse was joined to a normal mouse, the results were different.
The genetically obese mouse gained weight more slowly, and the paired animals survived for months.
Normal-normal control pairs also survived, although Coleman observed smaller fat pads than in normal mice that had not undergone parabiosis.
These results supported an explanation involving a circulating signal and the ability of different animals to respond to it.
According to Coleman’s interpretation, the ob/ob mice could respond to the signal but could not produce enough of it themselves.
The db/db mice produced the signal but were unable to respond to it, potentially because a receptor or another part of the receiving mechanism was defective.
Additional experiments involving the hypothalamus pointed toward the brain as an important site for detecting the signal.
The differences between the animal pairings gave Coleman a biological model that could explain why two distinct mutations produced a similar extreme-obesity phenotype.
One mutation appeared to prevent the body from sending the message. The other appeared to prevent it from receiving the message.
The experiments did not establish the chemical identity of the signal. They also could not measure food intake separately in every paired animal, a limitation later scientific reviews have emphasized.
Nevertheless, the overall pattern was sufficiently distinctive to provide a framework for searching for the underlying molecule.
The 1994 Discovery That Confirmed Coleman’s Prediction
Coleman had inferred a biological signaling pathway using experiments that preceded modern gene-sequencing techniques.
What he could not do was isolate and identify the circulating factor.
He continued searching, including investigating fatty acids and lipid extracts from adipose tissue, but those efforts did not reveal the molecule.
The definitive breakthrough arrived in 1994.
Jeffrey Friedman and colleagues at Rockefeller University published a landmark paper in Nature titled “Positional cloning of the mouse obese gene and its human homologue.”
The researchers identified the gene affected in ob/ob mice and showed that it encoded a protein hormone produced predominantly by adipose tissue.
The hormone was named leptin.
In 1995, Louis Tartaglia and colleagues identified a leptin receptor. Research published in 1996 then connected the db/db mutation to abnormalities in the leptin receptor gene.
The molecular findings explained why the two mouse strains behaved differently.
The ob/ob animals lacked functional leptin but retained the ability to respond to it.
The db/db animals had defective leptin signaling through the receptor, even though they could produce large quantities of the hormone.
Leptin was also shown to act through pathways in the hypothalamus, helping connect the body’s energy stores to brain circuits regulating feeding and metabolism.
The discovery changed how adipose tissue was understood.
Fat was no longer simply a passive energy reserve. It was an endocrine tissue capable of communicating information about stored energy to the brain.
The basic mechanism Coleman had inferred from his surgically paired mice was now identifiable at the level of genes, hormones and receptors.
More than 20 years after Coleman’s experiments, molecular biology finally identified the circulating hormone his mouse studies had predicted.
Recognition, Retirement and a Careful Message About Leptin
Coleman’s research initially faced considerable skepticism.
For much of the twentieth century, obesity was commonly framed primarily as a behavioral problem involving overeating and self-control.
His experiments suggested that inherited biological defects could substantially affect hunger, body weight and metabolism.
Although he received the Claude Bernard Award in 1977, broad recognition of the pathway developed more fully after leptin was identified.
Coleman retired from The Jackson Laboratory in 1993, a year before the molecular discovery that would vindicate much of his work.
He was elected to the United States National Academy of Sciences in 1998.
His later honors included the Gairdner International Award in 2005, the Shaw Prize in 2009 and the Albert Lasker Award for Basic Medical Research in 2010, which he shared with Jeffrey Friedman.
In 2013, he also received the King Faisal International Prize for Medicine and the BBVA Foundation Frontiers of Knowledge Award.
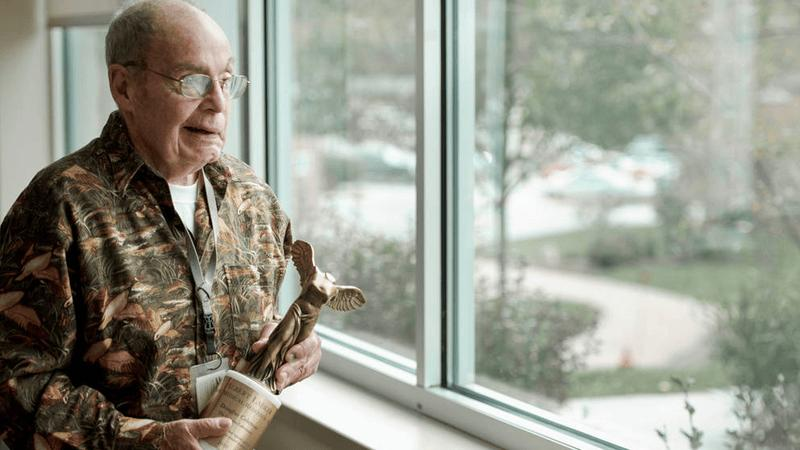
In a 2010 essay published in Nature Medicine for the Lasker Award, Coleman reflected on the long path from his early mouse experiments to the identification of leptin.
He also resisted the idea that the hormone would become a universal treatment for obesity.
Leptin replacement can produce dramatic benefits in people with rare inherited leptin deficiency, whose bodies cannot produce sufficient functional hormone.
But most people with common obesity do not lack leptin.
Many have elevated circulating leptin concentrations and reduced responsiveness to its effects, a phenomenon commonly described as leptin resistance.
Giving more hormone therefore does not reliably produce the same effect seen in people whose obesity is caused by leptin deficiency.
Coleman’s conclusion was characteristically cautious: understanding a hormone’s role in disease does not automatically make replacing that hormone a treatment for every form of the disease.
He died on April 16, 2014, at his home in Maine, at the age of 82.
His son Thomas described a scientist who had spent his entire professional life at a single laboratory and who, during retirement, became involved in conservation and local community activities.
The obituary also records that Coleman donated his scientific prize money to charitable causes.
Why Douglas Coleman’s Experiments Still Matter
Modern obesity medicine involves multiple hormonal pathways, including leptin, GLP-1, GIP and melanocortin signaling.
Drugs such as semaglutide and tirzepatide do not work by replacing leptin. They act through other hormone receptors involved in metabolic regulation and appetite.
Nevertheless, the broader conceptual foundation is related: body weight is influenced by biological signals linking peripheral organs to the brain.
Coleman’s experiments helped demonstrate that this signaling could be disrupted by specific inherited defects.
The leptin pathway itself eventually became relevant to targeted pharmaceutical development.
Rhythm Pharmaceuticals developed setmelanotide, sold as Imcivree, which acts on the melanocortin-4 receptor, a component of an appetite-regulating pathway downstream of leptin signaling.
The medicine has received regulatory approval for specific rare genetic and syndromic forms of obesity, rather than common obesity generally.
Grey Pepper has previously examined the company in its coverage of Rhythm Pharmaceuticals and the financing of Imcivree.
Leptin biology also remains relevant to research involving brain inflammation, immunity and metabolism.
One recent example is the experimental work on APOE4, microglia and leptin responses in mice, which explores how the brain’s immune environment may affect metabolic signaling.
None of those later developments means that Coleman single-handedly discovered all the biology underlying modern obesity treatment.
The history includes earlier physiological theories, other parabiosis researchers, Coleman’s collaborators, and the molecular biologists who eventually identified leptin and its receptor.
His particular contribution was experimentally distinguishing the absence of a circulating signal from the inability to respond to one.
It was a remarkably precise conclusion to draw without knowing the molecule’s name, its amino acid sequence or the identity of its receptor.
The experiments were performed entirely in mice, and the mutations involved represent unusual genetic causes of extreme obesity. They should not be treated as a complete explanation for common human obesity.
Nor was the full mechanism settled in the early papers. Individual food intake was difficult to measure in surgically paired animals, and several interpretations required additional experiments.
But the predictions proved sufficiently accurate to guide subsequent research for decades.
That is what makes the story enduring.
A scientist working in a mouse genetics laboratory used an old experimental technique to investigate a biological signal nobody could yet identify.
When molecular biology finally caught up, the missing hormone had a name: leptin.
Douglas Coleman did not need to see the molecule to understand that the body was sending a message. He needed two different mutations, a shared circulation and the patience to follow an unexpected result.
Sources and Further Reading
- Coleman TR. Douglas L. Coleman, 1931–2014. Diabetologia. Published October 7, 2014. DOI: 10.1007/s00125-014-3393-7.
- Coleman DL. A historical perspective on leptin. Nature Medicine. October 2010. Lasker Basic Medical Research Award commentary.
- Coleman DL, Hummel KP. Effects of parabiosis of normal with genetically diabetic mice. American Journal of Physiology. 1969;217(5):1298–1304. DOI: 10.1152/ajplegacy.1969.217.5.1298.
- Coleman DL. Effects of parabiosis of obese with diabetes and normal mice. Diabetologia. 1973;9(4):294–298. DOI: 10.1007/BF01221857.
- Zhang Y, Proenca R, Maffei M, Barone M, Leopold L, Friedman JM. Positional cloning of the mouse obese gene and its human homologue. Nature. 1994;372:425–432. DOI: 10.1038/372425a0.
- Society for the Study of Ingestive Behavior. Douglas Coleman’s Parabiosis Studies in Obese and Diabetic Mice. Historical scientific commentary by Ruth Harris.
- US Food and Drug Administration. Imcivree (setmelanotide) Prescribing Information, 2026.